
Research article · 2024 · Full text
International Journal of Advanced Chemistry ResearchCalculating theoretical octane numbers of various petroleum derivatives using quantum mechanical DFT methods
Volume 6(2) · Pages 16–21 · DOI: 10.33545/26646781.2024.v6.i2a.220 ↗
Research areasThermodynamic properties of mixturesFree Radicals and AntioxidantsVarious Chemistry Research Topics
Abstract
Author-writtenThis research included calculating the octane number of a group of chemical compounds using the (DFT) method by using multiple regression analysis of the relationship between the practical octane number values with the theoretically calculated physical variables, where a number of variables were calculated including the calculation of energy functions, including the highest occupied molecular orbital (HOMO) and the lowest unoccupied molecular orbital (LOMO), hardness (η), chemical potential (µ), and spherical electrophilic index (W), and the thermodynamic functions (∆G, ∆H, ∆S, ∆E) and dipole moment were calculated. They were evaluated from the correlation coefficient relationship (R2). Through the four-way statistical analysis, we obtained the best correlation coefficient for the variables calculated using the DFT method, where (R2) showed the highest values for the following variables: (∆E+HOMO+∆G+µ) (R2=0.793) (∆S+ LOMO + ∆G +W) (R2=0.753). Through the single and multiple statistical analysis, the most influential variable on the octane number values is (HOMO).
Summary
Source-checkedPlain-language reading of the full paper, prepared with AI and checked against the text. Each point links to the authors' own words.
Why it matters
- The octane number is used as a measure of the ignition quality or ability of gasoline. View evidence ↗
- Computational chemistry offers a safer and less expensive alternative to experimental methods because it avoids the risks and costs associated with using chemicals. View evidence ↗
- Density Function Theory (DFT) is considered a highly accurate and fast calculation method for finding electronic properties of molecules in a stable state. View evidence ↗
The question
- The study aimed to calculate the octane number of various chemical compounds using the DFT method and multiple regression analysis to relate theoretical physical variables to practical octane values. View evidence ↗
What they did
- The research analyzed nine petroleum derivatives, including benzene, toluene, ethyl benzene, propyl benzene, butyl benzene, o-xylene, m-xylene, p-xylene, and mesitylene. View evidence ↗
- The authors used Gaussian 09 software and DFT methods to calculate dipole moments, thermodynamic functions (∆H, ∆G, ∆S, ∆E), orbital energies (HOMO, LOMO), and derived energy functions (hardness, chemical electron potential, and spherical electrophilic index). View evidence ↗
- SPSS software was used to perform the statistical analysis of the calculated variables. View evidence ↗
What they found
- Statistical analysis identified the Highest Occupied Molecular Orbital (HOMO) as the most influential variable affecting octane number values. View evidence ↗
- Model 5 was determined to be the most effective for calculating theoretical octane numbers, achieving the highest correlation coefficient (R2) of 0.793. View evidence ↗
- Model 5 showed a strong correlation (R = 0.890), explaining 79.3% of the variance in octane numbers using the independent variables. View evidence ↗
- The study derived a statistical equation for theoretical octane number based on ∆E, HOMO, ∆G, and µ. View evidence ↗
- Theoretical octane numbers were compared to true values, showing exact matches for benzene (102) and propyl benzene (110), with the largest difference of 20 units observed for o-xylene. View evidence ↗
What it means
- The authors conclude that the proposed variables are strong predictors for calculating octane numbers in gasoline derivatives, expanding the potential of computational chemistry. View evidence ↗
Visual summary
3 visualsIllustrations explain the paper; charts are replotted from the paper's own table values.
DFT-to-Octane Prediction Workflow
AI-generated conceptual illustration (petroleum derivative molecules). It explains the idea and does not show study data.
View source evidence ↗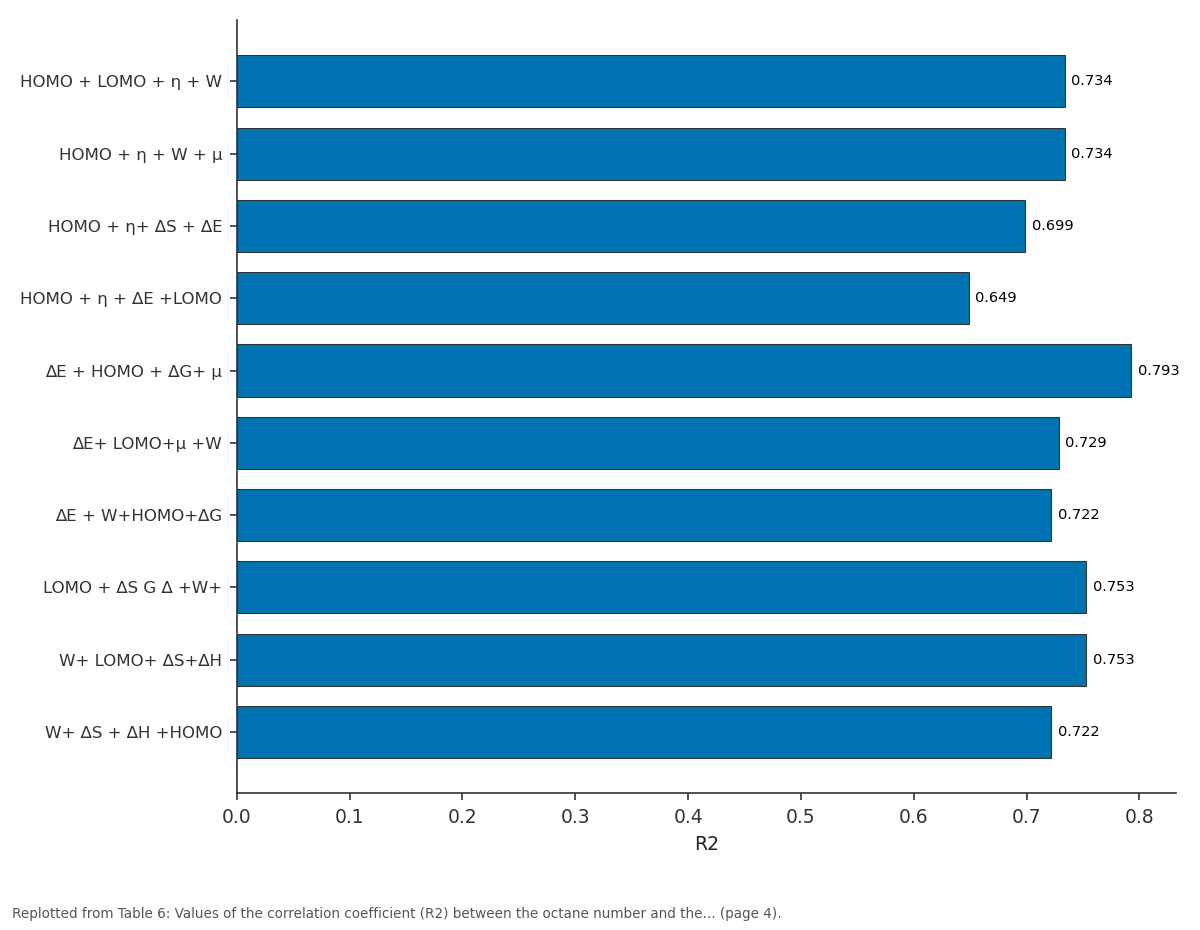
R2: HOMO + LOMO + η + W vs HOMO + η + W + µ vs HOMO + η+ ∆S + ∆E vs HOMO + η + ∆E +LOMO vs ∆E + HOMO + ∆G+ µ
Replotted from Table 6 (page 4) of the original paper. Every bar is the paper's own table value.
View source evidence ↗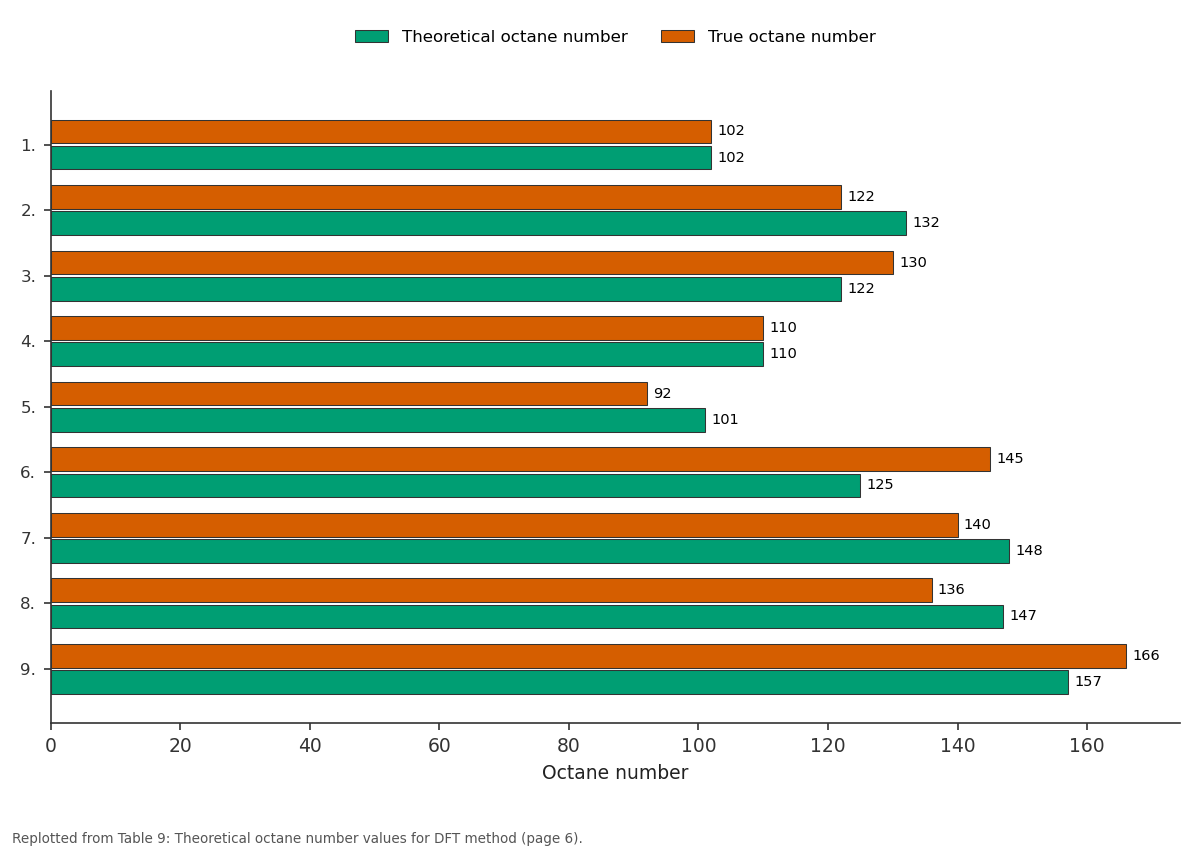
Theoretical octane number and True octane number: 1. vs 2. vs 3. vs 4. vs 5.
Replotted from Table 9 (page 6) of the original paper. Every bar is the paper's own table value.
View source evidence ↗Figures & tables
From the original paper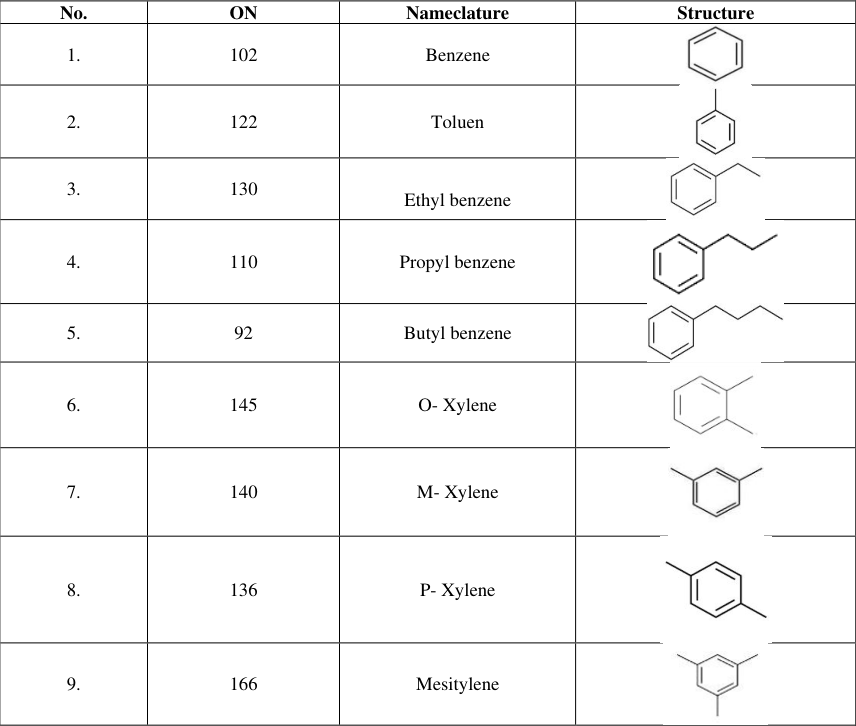 Table 1
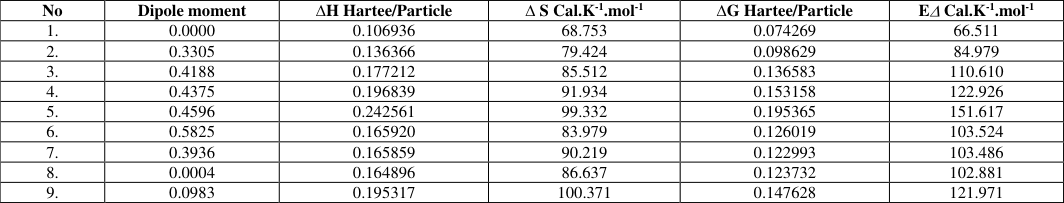 Table 2
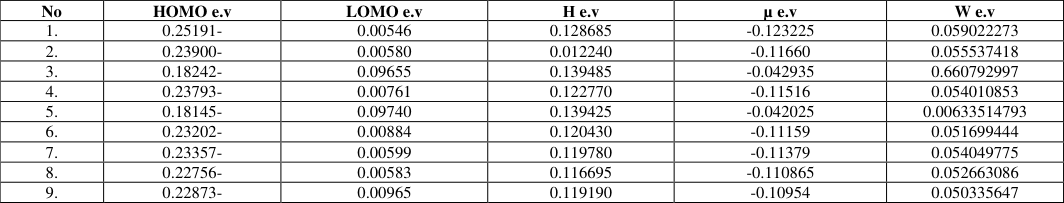 Table 3
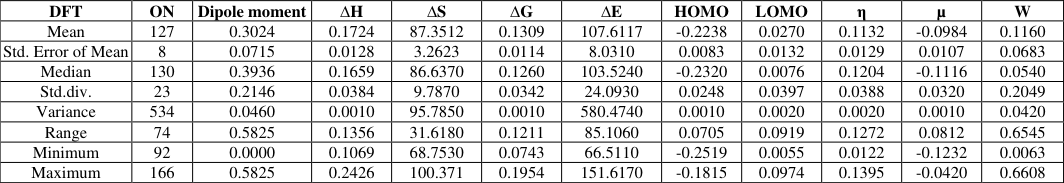 Table 4
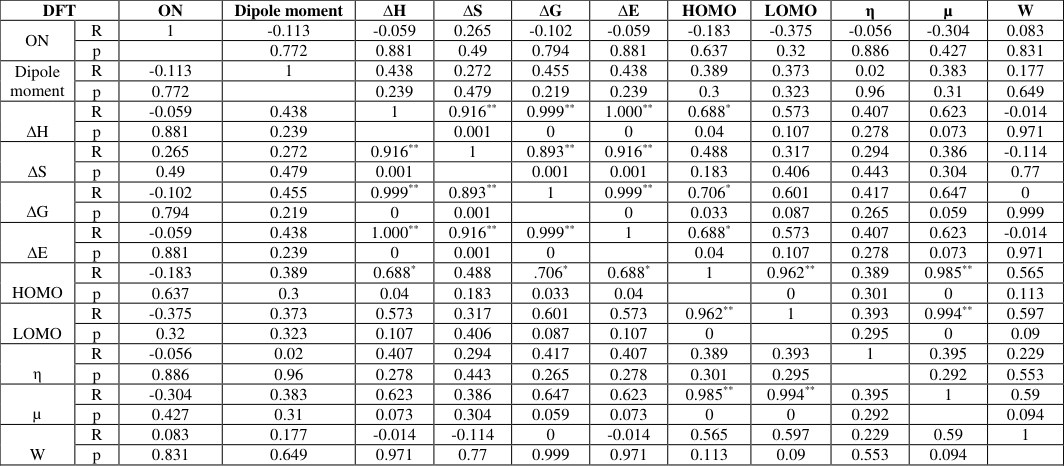 Table 5
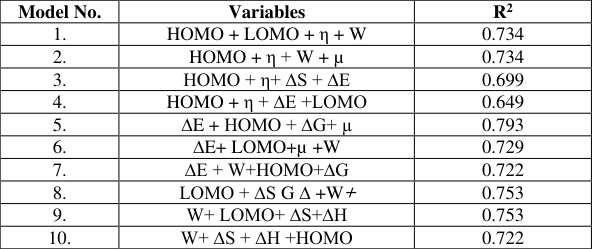 Table 6
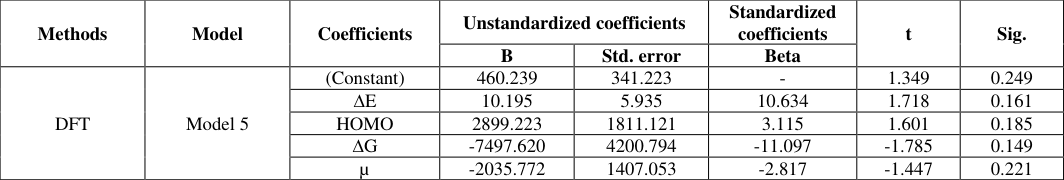 Table 7
 Table 8
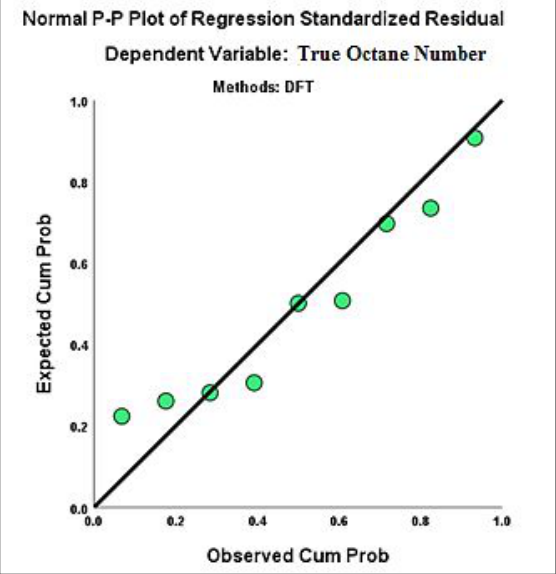 Figure 1
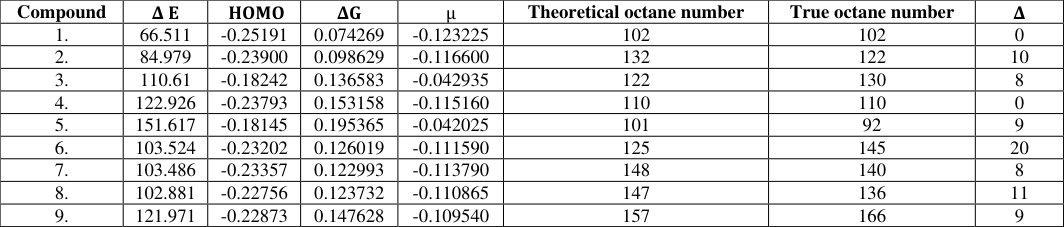 Table 9
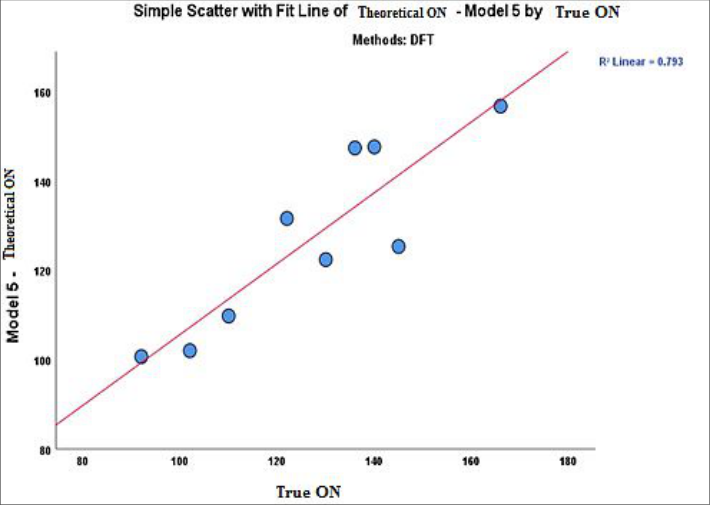 Figure 2
Introduction Octane Number
It is a measure of the ignition quality or ignition ability of gasoline. Research Octane Number (RON) refers to normal road performance under moderate inlet temperature (65.6 °C) and rpm (600 rpm). Engine octane number (MON) refers to high-speed performance under severe inlet temperatures (148.9 °C) and 900 rpm. Research Octane Number (RON) and Engine Octane Number (MON) are evaluated using ASTM D2699 and ASTM D2700, respectively [1,2] .
Computational chemistry
Computational chemistry is one of the branches of physical chemistry, including quantum mechanics and molecular mechanics. Its purpose is to find the most important properties of chemical compounds and compare them with practical values. It is one of the main branches of research used in the analysis, diagnosis, and support of scientific research [3] . Computational chemistry depends on the use of computers instead of chemicals and the creation of effective mathematical methods to solve theoretical problems through advanced computational methods. John People is considered the first to introduce computers into quantum chemistry in the fifties of the last century by creating new integrated algorithms, highly efficient in calculations, and then marketing computational programs such as the Gaussian program [4] . Computational chemistry is characterized by the fact that its calculations are easy to perform compared to In experiments that are often difficult to perform and involve the risks of chemicals as well as less expensive because they do not require chemicals, and can be performed on any system even if it is not present, while experiments are limited to relatively stable molecules, however, computational chemistry is not an alternative to experimental methods, but it plays an important role in knowing the properties of chemical compounds [5] .
The term computational chemistry is used when the mathematical method has been developed sufficiently to be applied using computer programs [6] .
Density Function Theory (DFT)
The first to use and develop this theory were the researchers (Hohenberge-Kohn) in 1964 AD, and it later became one of the important components related to the basic analysis methods that were used extensively in theoretical studies, especially after the development in the field of electronic computers. This theory is based on finding the electronic properties of the molecule in the stable state, which are found using the electronic density. The advantage of this theory is that it includes finding the approximate electronic correlations with the same The accuracy calculated by the well-known method (Hartrey-Fock) (HF) is usually faster than the experimental method and is considered the most accurate method in calculations, so it can be used as one of the basic calculation methods (Ab-initio). It is similar to the basic calculations and semi-empirical calculations in terms of its reliance on the Schrodinger equation, but it differs from the two methods in that it does not calculate a wave function but derives the distribution of electrons (electron density function) directly, where the electron density is expressed as the square of the function. Wave, the basis of this method is to simulate the actual distribution of the electron density of the system, so the larger the set of functions used to model the electrons, the more accurate the calculations become, and the results are much better, as they have been developed by introducing electronic exchange and correlation functions that increase the accuracy of the calculations. There are many approximations used in this method, including: LDAM: Local Density Approximation. LSDA: Local spin Density Approximation. GCM: Gradient Corrected Method.
Hybrid functional function
In this, the functional function theory (DFT) is combined with the Hartree-Fock theory (HF) to describe the functional function of the exchange energy, including: B3LYP: Becke's 3 parameter exchange correlation functional, which uses 3 Parameters and LYP, the Lee, Yang and Parr correlation functional.
This function is considered the most used in calculations due to the accuracy of the results that can be obtained. However, new functions are constantly being developed to increase the accuracy of calculations [7,8] .
Theoretical calculation
Quantum mechanics methods were used in this study, represented by the DFT method, by using these theoretical methods to find the values of the octane number by the (Chem. Office) program [9] . The calculations were performed using the following methods 1. Ab-initio methods (DFT).
Gaussian 09 program
Steps of theoretical calculations for the compounds under study using Gaussian 09: 1. Drawing the molecular formula of the compounds using the (Chem. Office) program. 2. Transferring the compound drawn in the (Chem.
Office) program to the (Chem. 3D) program. 3. Save the compound in the (Chem. 3D) program in the file format (Input File). 4. Open the program (Gaussian 0.6) then open the file saved in (Input File) format. 5. Select the (Calculate) button, then select (Gaussian Calculation Setup). 6. From the Job Type button, select (Optimization), then select (Minimum) to obtain the lowest energy, which will be more stable and have the best spatial shape. 7. After that, from the (Method) button, select the theoretical method and determine the basis set. 8. After that, select (Submit) to perform the calculations in the (Gaussian 09W) program. 9. After finishing and completing the results, they will be saved via (Notepad). 10. The statistical analysis is performed using the SPSS program.
Statistical analysis
The statistical analysis was conducted between the selected and calculated variables (HOMO, LOMO, dipole moment, hardness, electronic potential, spherical electrophilic index, ∆H, ∆G, ∆S, ∆Ε) using theoretical methods with the practical Octane Number (ON) values calculated using SPSS
Program
The correlation coefficient (R 2 ) values were adopted as a reference for this purpose to know the strength and type of relationship between the calculated variables and the practical Octane Number (ON) values, as the statistical analysis was conducted between the variables with each other and with the practical Octane Number (ON) values using the methods used, where each method was taken separately and with the following steps:
1. The calculated variables of the studied compounds (HOMO, LOMO, dipole moment, Hardness, electron potential, spherical electrophilic index, ∆H, ∆G, ∆S, ∆Ε) were entered as non-dependent variables with the practical octane number values that are dependent variables.
2. The variables were analyzed through a series of steps in the SPSS program using the following steps: (Linear + regression + analysis). 3. The values of the correlation coefficient (R 2 ) are then obtained, since knowing the values of (R 2 ) determines the variables that have a significant effect on the Octane Number (ON) in the simple regression analysis. 4. We then conduct the process of multiple statistical analysis to determine the strength of the relationship between the variables that were used with each other and with the scientific octane number values obtained from the literature.
Theoretical calculations
In this research, the compounds were studied theoretically and the Octane Number (ON) values were extracted theoretically, as a set of variables were calculated, including the thermodynamic functions (∆H, ∆G, ∆S, ∆Ε), dipole moment and orbital energy (HOMO, LOMO), and energy functions (hardness η, electron-chemical potential µ, spherical electrophilic index W). The octane number values were calculated for a number of compounds to obtain the best relationship between the variables and the octane number values. (Table 4) shows the descriptive statistics of the variables studied using the DFT method. The table shows that the average octane number of the studied samples is 127, and the average polar moment of the studied materials is about 0.3024. The enthalpy (∆H), entropy (∆S), and free energy (∆G) changes are around the average values of 87.3512, 0.1724, and 0.1309 respectively, while the average energy change (∆E) of the studied materials is about 107.6117. For the highest occupied molecular orbital (HOMO) and the lowest unoccupied molecular orbital (LOMO), the average values are -0.2238 and 0.0270, respectively. The average transport efficiency (η) of the studied materials is 0.1132, and the average polarity (µ) is -0.0984. It is noted from the attached (Table 9) and from the results of the regression analysis, that there is a significant correlation between the practical octane number values obtained from the literature with the theoretically calculated octane number values, with some differences in some compounds for the four methods used. This is evidence of the strength of the proposed variables for calculating the octane number values for some gasoline derivatives [10] . This study opens up broad horizons in the field of software chemistry (computational chemistry).
| Compound | 𝚫 𝐄 | 𝐇𝐎𝐌𝐎 | 𝚫𝐆 | µ | Theoretical octane number | True octane number | 𝚫 |
|---|---|---|---|---|---|---|---|
| 1. | 66.511 | -0.25191 | 0.074269 | -0.123225 | 102 | 102 | 0 |
| 2. | 84.979 | -0.23900 | 0.098629 | -0.116600 | 132 | 122 | 10 |
| 3. | 110.61 | -0.18242 | 0.136583 | -0.042935 | 122 | 130 | 8 |
| 4. | 122.926 | -0.23793 | 0.153158 | -0.115160 | 110 | 110 | 0 |
| 5. | 151.617 | -0.18145 | 0.195365 | -0.042025 | 101 | 92 | 9 |
| 6. | 103.524 | -0.23202 | 0.126019 | -0.111590 | 125 | 145 | 20 |
| 7. | 103.486 | -0.23357 | 0.122993 | -0.113790 | 148 | 140 | 8 |
| 8. | 102.881 | -0.22756 | 0.123732 | -0.110865 | 147 | 136 | 11 |
| 9. | 121.971 | -0.22873 | 0.147628 | -0.109540 | 157 | 166 | 9 |
Table text extracted from the original PDF; superscript letters are shown inline. The original table image is below.
View original table image
| DFT | ON | Dipole moment | ∆H | ∆S | ∆G | ∆E | HOMO | LOMO | η | µ | W |
|---|---|---|---|---|---|---|---|---|---|---|---|
| Mean | 127 | 0.3024 | 0.1724 | 87.3512 | 0.1309 | 107.6117 | -0.2238 | 0.0270 | 0.1132 | -0.0984 | 0.1160 |
| Std. Error of Mean | 8 | 0.0715 | 0.0128 | 3.2623 | 0.0114 | 8.0310 | 0.0083 | 0.0132 | 0.0129 | 0.0107 | 0.0683 |
| Median | 130 | 0.3936 | 0.1659 | 86.6370 | 0.1260 | 103.5240 | -0.2320 | 0.0076 | 0.1204 | -0.1116 | 0.0540 |
| Std.div. | 23 | 0.2146 | 0.0384 | 9.7870 | 0.0342 | 24.0930 | 0.0248 | 0.0397 | 0.0388 | 0.0320 | 0.2049 |
| Variance | 534 | 0.0460 | 0.0010 | 95.7850 | 0.0010 | 580.4740 | 0.0010 | 0.0020 | 0.0020 | 0.0010 | 0.0420 |
| Range | 74 | 0.5825 | 0.1356 | 31.6180 | 0.1211 | 85.1060 | 0.0705 | 0.0919 | 0.1272 | 0.0812 | 0.6545 |
| Minimum | 92 | 0.0000 | 0.1069 | 68.7530 | 0.0743 | 66.5110 | -0.2519 | 0.0055 | 0.0122 | -0.1232 | 0.0063 |
| Maximum | 166 | 0.5825 | 0.2426 | 100.371 | 0.1954 | 151.6170 | -0.1815 | 0.0974 | 0.1395 | -0.0420 | 0.6608 |
Table text extracted from the original PDF; superscript letters are shown inline. The original table image is below.
View original table image
Figures and tables from the paper
| No. | ON | Nameclature | Structure |
|---|---|---|---|
| 1. | 102 | Benzene | |
| 2. | 122 | Toluen | |
| 3. | 130 | Ethyl benzene | |
| 4. | 110 | Propyl benzene | |
| 5. | 92 | Butyl benzene | |
| 6. | 145 | O- Xylene | |
| 7. | 140 | M- Xylene | |
| 8. | 136 | P- Xylene | |
| 9. | 166 | Mesitylene |
Table text extracted from the original PDF; superscript letters are shown inline. The original table image is below.
View original table image
| No | Dipole moment | ∆H Hartee/Particle | ∆ S Cal.K-1.mol-1 | ∆G Hartee/Particle | E∆ Cal.K-1.mol-1 |
|---|---|---|---|---|---|
| 1. | 0.0000 | 0.106936 | 68.753 | 0.074269 | 66.511 |
| 2. | 0.3305 | 0.136366 | 79.424 | 0.098629 | 84.979 |
| 3. | 0.4188 | 0.177212 | 85.512 | 0.136583 | 110.610 |
| 4. | 0.4375 | 0.196839 | 91.934 | 0.153158 | 122.926 |
| 5. | 0.4596 | 0.242561 | 99.332 | 0.195365 | 151.617 |
| 6. | 0.5825 | 0.165920 | 83.979 | 0.126019 | 103.524 |
| 7. | 0.3936 | 0.165859 | 90.219 | 0.122993 | 103.486 |
| 8. | 0.0004 | 0.164896 | 86.637 | 0.123732 | 102.881 |
| 9. | 0.0983 | 0.195317 | 100.371 | 0.147628 | 121.971 |
Table text extracted from the original PDF; superscript letters are shown inline. The original table image is below.
View original table image
| No | HOMO e.v | LOMO e.v | Η e.v | µ e.v | W e.v |
|---|---|---|---|---|---|
| 1. | 0.25191- | 0.00546 | 0.128685 | -0.123225 | 0.059022273 |
| 2. | 0.23900- | 0.00580 | 0.012240 | -0.11660 | 0.055537418 |
| 3. | 0.18242- | 0.09655 | 0.139485 | -0.042935 | 0.660792997 |
| 4. | 0.23793- | 0.00761 | 0.122770 | -0.11516 | 0.054010853 |
| 5. | 0.18145- | 0.09740 | 0.139425 | -0.042025 | 0.00633514793 |
| 6. | 0.23202- | 0.00884 | 0.120430 | -0.11159 | 0.051699444 |
| 7. | 0.23357- | 0.00599 | 0.119780 | -0.11379 | 0.054049775 |
| 8. | 0.22756- | 0.00583 | 0.116695 | -0.110865 | 0.052663086 |
| 9. | 0.22873- | 0.00965 | 0.119190 | -0.10954 | 0.050335647 |
Table text extracted from the original PDF; superscript letters are shown inline. The original table image is below.
View original table image
| DFT | — | ON | Dipole moment | ∆H | ∆S | ∆G | ∆E | HOMO | LOMO | η | µ | W |
|---|---|---|---|---|---|---|---|---|---|---|---|---|
| ON | R | 1 | -0.113 | -0.059 | 0.265 | -0.102 | -0.059 | -0.183 | -0.375 | -0.056 | -0.304 | 0.083 |
| p | 0.772 | 0.881 | 0.49 | 0.794 | 0.881 | 0.637 | 0.32 | 0.886 | 0.427 | 0.831 | ||
| Dipole moment | R | -0.113 | 1 | 0.438 | 0.272 | 0.455 | 0.438 | 0.389 | 0.373 | 0.02 | 0.383 | 0.177 |
| p | 0.772 | 0.239 | 0.479 | 0.219 | 0.239 | 0.3 | 0.323 | 0.96 | 0.31 | 0.649 | ||
| ∆H | R | -0.059 | 0.438 | 1 | 0.916** | 0.999** | 1.000** | 0.688* | 0.573 | 0.407 | 0.623 | -0.014 |
| p | 0.881 | 0.239 | 0.001 | 0 | 0 | 0.04 | 0.107 | 0.278 | 0.073 | 0.971 | ||
| ∆S | R | 0.265 | 0.272 | 0.916** | 1 | 0.893** | 0.916** | 0.488 | 0.317 | 0.294 | 0.386 | -0.114 |
| p | 0.49 | 0.479 | 0.001 | 0.001 | 0.001 | 0.183 | 0.406 | 0.443 | 0.304 | 0.77 | ||
| ∆G | R | -0.102 | 0.455 | 0.999** | 0.893** | 1 | 0.999** | 0.706* | 0.601 | 0.417 | 0.647 | 0 |
| p | 0.794 | 0.219 | 0 | 0.001 | 0 | 0.033 | 0.087 | 0.265 | 0.059 | 0.999 | ||
| ∆E | R | -0.059 | 0.438 | 1.000** | 0.916** | 0.999** | 1 | 0.688* | 0.573 | 0.407 | 0.623 | -0.014 |
| p | 0.881 | 0.239 | 0 | 0.001 | 0 | 0.04 | 0.107 | 0.278 | 0.073 | 0.971 | ||
| HOMO | R | -0.183 | 0.389 | 0.688* | 0.488 | .706* | 0.688* | 1 | 0.962** | 0.389 | 0.985** | 0.565 |
| p | 0.637 | 0.3 | 0.04 | 0.183 | 0.033 | 0.04 | 0 | 0.301 | 0 | 0.113 | ||
| LOMO | R | -0.375 | 0.373 | 0.573 | 0.317 | 0.601 | 0.573 | 0.962** | 1 | 0.393 | 0.994** | 0.597 |
| p | 0.32 | 0.323 | 0.107 | 0.406 | 0.087 | 0.107 | 0 | 0.295 | 0 | 0.09 | ||
| η | R | -0.056 | 0.02 | 0.407 | 0.294 | 0.417 | 0.407 | 0.389 | 0.393 | 1 | 0.395 | 0.229 |
| p | 0.886 | 0.96 | 0.278 | 0.443 | 0.265 | 0.278 | 0.301 | 0.295 | 0.292 | 0.553 | ||
| µ | R | -0.304 | 0.383 | 0.623 | 0.386 | 0.647 | 0.623 | 0.985** | 0.994** | 0.395 | 1 | 0.59 |
| p | 0.427 | 0.31 | 0.073 | 0.304 | 0.059 | 0.073 | 0 | 0 | 0.292 | 0.094 | ||
| W | R | 0.083 | 0.177 | -0.014 | -0.114 | 0 | -0.014 | 0.565 | 0.597 | 0.229 | 0.59 | 1 |
| p | 0.831 | 0.649 | 0.971 | 0.77 | 0.999 | 0.971 | 0.113 | 0.09 | 0.553 | 0.094 |
Table text extracted from the original PDF; superscript letters are shown inline. The original table image is below.
View original table image
| Model No. | Variables | R2 |
|---|---|---|
| 1. | HOMO + LOMO + η + W | 0.734 |
| 2. | HOMO + η + W + µ | 0.734 |
| 3. | HOMO + η+ ∆S + ∆E | 0.699 |
| 4. | HOMO + η + ∆E +LOMO | 0.649 |
| 5. | ∆E + HOMO + ∆G+ µ | 0.793 |
| 6. | ∆E+ LOMO+µ +W | 0.729 |
| 7. | ∆E + W+HOMO+∆G | 0.722 |
| 8. | LOMO + ∆S G ∆ +W+ | 0.753 |
| 9. | W+ LOMO+ ∆S+∆H | 0.753 |
| 10. | W+ ∆S + ∆H +HOMO | 0.722 |
Table text extracted from the original PDF; superscript letters are shown inline. The original table image is below.
View original table image
| Methods | Model | Coefficients | Unstandardized coefficients | — | Standardized coefficients | t | Sig. |
|---|---|---|---|---|---|---|---|
| B | Std. error | Beta | |||||
| DFT | Model 5 | (Constant) | 460.239 | 341.223 | - | 1.349 | 0.249 |
| ∆E | 10.195 | 5.935 | 10.634 | 1.718 | 0.161 | ||
| HOMO | 2899.223 | 1811.121 | 3.115 | 1.601 | 0.185 | ||
| ∆G | -7497.620 | 4200.794 | -11.097 | -1.785 | 0.149 | ||
| µ | -2035.772 | 1407.053 | -2.817 | -1.447 | 0.221 |
Table text extracted from the original PDF; superscript letters are shown inline. The original table image is below.
View original table image
| Model summary | — | — | — | — | — |
|---|---|---|---|---|---|
| Methods | Model | R | R square | Adjusted R square | Std. error of the estimate |
| DFT | Model 5 | 0.890 | 0.793 | 0.585 | 14.877 |
Table text extracted from the original PDF; superscript letters are shown inline. The original table image is below.
View original table image
References
4 in the original paper- Al-Abady FM. Quantum-chemical calculation methods to determine the ionization constants of N-substituted amino acid derivatives. Eur. Chem. Bull. 2017;6(8):343-349.
- Van Houten J. A century of chemical dynamics traced through the Nobel prizes: 1998: Walter Kohn and John Pople. J Chem Educ. 2002;79(11):1297.
- Kubicki JD, Ohno T. Integrating density functional theory modeling with experimental data to understand and predict sorption reactions: Exchange of salicylate for phosphate on goethite. Soil Syst. 2020;4(2):27.
- Kerwin SM. Chem Bio Office Ultra 2010 Suite; c2010.